Over 30 years of reporting on Congress, federal agencies and the White House for corporate America as well as national trade and professional associations.

Controlling Drug Prices

Human Resource Executive magazine online

January 22, 2007


Congress is offering several variations on ways for Medicare to lower drug prices -- a process that may reverberate in the corporate sector.

By Stephen Barlas

An attempt by House Democrats aimed at forcing drug manufacturers to lower prices charged to Medicare could well have an impact on corporate health-insurance spending as well.

Though the bill, which passed the House on Jan. 12, requires the federal government to negotiate directly with drug companies, odds are it will not pass Congress. Nevertheless, it may pave the way for a federal role that perhaps could include a new requirement for drug price "transparency."

Any downward pressure on Medicare drug prices would reverberate in the corporate sector, says Cara Jareb, director of retiree medical consulting for Watson Wyatt Worldwide, which is headquartered in Bethesda, Md.

She cites statistics from the Center for Medicare and Medicaid Services, the agency which administers Medicare, showing that of the 38 million Medicare participants in Part D, 6.9 million are corporate retirees in corporate plans for whom each company gets a subsidy from Medicare.

Another 16.4 million Part D participants actually get their drug benefits from one of the Medicare plans, but those individuals also get payments from their former employers to underwrite their Part D participation.

If drug companies are forced to cut their prices to Medicare, they are likely to raise them to private payers, a phenomenon called "cost shifting."

Steve Wojcik, vice president of public policy at the Washington-based National Business Group on Health, which represents Fortune 500 companies, says, "There is a legitimate concern that cost shifting could occur."

Senate leaders from both parties have all but declared the House bill dead on arrival, and even if it somehow passed the Senate, President Bush has threatened to veto it. J.D. Piro, a principal at Lincolnshire, Ill.-based Hewitt Associates, says the House does not have the votes to override a veto.

That bill aside, however, lowering Medicare drug costs remains a potent political issue.

Jareb points to a recent Kaiser Permanente survey which showed 85 percent of people questioned said Medicare can get lower prices from drug companies than it currently obtains.

Given that political pressure, the Senate may opt to pass a more limited approach, such as the one taken by Sens. Ron Wyden, D.-Ore., and Olympia Snowe, R.-Maine. Their bill requires the secretary of Health and Human Services to negotiate prices when there is one brand-name drug in a therapeutic category or when a drug was created with substantial taxpayer funding for its research and development.

Another approach was suggested by Gerard Anderson, professor of health policy and management at Johns Hopkins University, who testified at hearings in the Senate Finance Committee on Jan. 11.

He wants drug-price transparency. "Unfortunately we do not know the prices that the Part D plans are paying for individual drugs," said Anderson, noting that the Center for Medicare and Medicaid Services "collects the data on prices, price concessions, rebates, and discounts but is prohibited from sharing this data or even analyzing it internally."

The Bush administration has pushed price transparency in tandem with its advocacy for health savings accounts, which give an individual tax advantages in conjunction with high-deductible health plans.

So the White House might find it politically difficult to oppose transparency in Medicare drug pricing.

"That is certainly a possibility if the Bush administration felt it had to compromise," says Piro.

But Mark Merritt, president of the industry trade group, Pharmaceutical Care Management Association, says if the prices PBMs pay drug companies for individual drugs become public, "it would be like playing poker with our cards face up."

Congress opens the SarbOx box

Financial Week

House legislation could go beyond federal regulators’ proposed relief

By Stephen Barlas

Not content waiting for the Securities and Exchange Commission to revise Sarbanes-Oxley, some Congressmen are taking things into their own hands by preparing to rewrite the law—opening opportunities for change and even lobbyist mischief.

Two House members are readying a version of a Sarbanes-Oxley reform bill as business groups consider whether proposed changes to Section 404 of that law, announced in December, go far enough.

Rep. Gregory Meeks, D-N.Y., and Rep. Tom Feeney, R-Fla., two mid-level members of the House Financial Services Committee, are making substantial changes to the Compete Act, a bill they introduced last year that included provisions to revise Sarbanes-Oxley.

Their bill will prove a test for the new Democratic House leadership, whose two kingpins made promises going into and following the 2006 congressional elections that they would pursue legislative changes to Sarbanes-Oxley.

Jameel Johnson, Mr. Meeks’ chief of staff, said he has been working with groups such as the Institute of Management Accountants, the Financial Services Roundtable and the Biotechnology Industry Organization on the bill’s provisions.

In 2006, BIO helped organize a coalition that sought significant changes to Sarbanes-Oxley. Many of those changes are absent from the interpretive guidance on Section 404 that the Securities Exchange Commission proposed in December and the Public Company Accounting Oversight Board’s proposed revisions to its auditing standard for 404.

The BIO coalition, which includes semiconductor, telecommunications and medical-device trade associations, wants smaller companies to be able to test their financial controls every second or third year, not annually as SarbOx now requires. The Meeks-Feeney bill will include such a provision that applies to companies of all sizes.

Mr. Johnson said the 2007 version of the bill would combine new provisions with some of those included in the bill when it was originally introduced on May 17, 2006. Sen. Jim DeMint, R-S.C., introduced the bill in the Senate at the same time in 2006.

He will reintroduce the bill in 2007, but details haven’t been worked out, according to a spokesman.

“Obviously, their efforts are well placed, and we want to encourage them to move forward,” said Alan Eisenberg, executive vice president of capital formation and business development at BIO.

Political pressure from high-tech companies led Rep. Rahm Emanuel, D-Ill., chair of the House Democratic caucus and ramrod of the Democratic Congressional Campaign Committee, to pledge at an off-the-record meeting of a major business group last September that the Democrats would “relegislate” SarbOx if they gained control of the House, according to someone who attended that meeting.

Then House Speaker Nancy Pelosi, D-Calif., made Mr. Emanuel’s promise public when she unveiled the Democrats’ “Innovation Agenda” at the National Press Club on Nov. 16. That agenda includes a proposal to require specifically tailored guidelines for small public companies to ensure Sarbanes-Oxley requirements are not overly burdensome.

Ms. Pelosi underscored that commitment the next day at a meeting with companies that are members of the American Electronics Association.

A Pelosi spokesman said she wants to see the public comments about the SEC and PCAOB proposals before deciding on a next step. A spokesman for Mr. Emanuel did not return phone calls or e-mails seeking comment.

Over a period of seven days beginning about a month after Ms. Pelosi put forth her party’s agenda, the SEC and PCAOB unveiled their packages of changes to SarbOx rules that were meant to dampen efforts by high-tech groups and the U.S. Chamber of Commerce to force the Democrats’ hands.

The PCAOB then proposed pulling Auditing Standard 2 and substituting a new AS5. PCAOB Chairman Mark Olson presumably echoed Ms. Pelosi by referring to his effort to “scale” audits to the “size and complexity of each company.”

But the Meeks-Feeney bill would go far beyond the SEC and PCAOB changes, which business groups have been busy analyzing.

Mr. Johnson said the 2007 bill would allow companies of all sizes to test their internal controls just once every two or three years after getting a good grade from auditors on their initial accounting review.

“Auditors who come in to test those companies’ financial controls as part of the attestation required by SOX should be able to rely on the analysis done by the federal regulators,” explained Mr. Johnson.

“The outside auditors should not have to test the controls again, as long as the company makes the federal or state review available to the outside auditor.”

This year’s bill may also include some provisions from the original bill, one of which exempted small businesses with a total capitalization under $700 million or product revenue under $125 million from complying with Section 404.

Another provision would allow small companies that hire internal consultants to do the management report and testing to talk with the company’s outside auditors before doing the testing so as to better focus their efforts. That provision will reappear in 2007, said Mr. Johnson. FW

Business Awaits Labor Dept. Rule on Default Pension Investments

December 2006 Financial Week

Some corporate pension fund fiduciaries will soon know how the Roman mythological god Janus felt. Treasurers and financial officers will be looking backward and forward simultaneously when the Department of Labor issues its final rule on default investments for automatic enrollment 401(k) plans. Mutual funds and investment firms will storm the front door offering investments. But trial lawyers may just as aggressively pursue fiduciaries through the back door.

Those changes, which corporate America hoped the DOL would publish by January 1, in time for the earliest company announcements on changes for 2007 plan years, implement a provision in the Pension Protection Act (PPA) of 2006, which President Bush signed last August. That law was the most significant change in pension law since the passage of the Employee Retirement Income Security Act (ERISA). The proposed rule issued late in September, the first of many expected to originate with the PPA, listed the types of investment vehicles companies could use as defaults for individual 401 (k)s in automatic enrollment plans.

However, many companies are worried that the final rule will expose them to past liability. That is because companies who now offer automatic enrollment heavily favor, as default investments, stable value and money market funds. Those plans could become legally problematic if the DOL excludes them as QDIAs, as it did in the proposed rule, injecting them with a whiff of imprudence.

Judy Schub, managing director of the Committee on Investment of Employee Benefits, says that she has heard considerable consternation from corporate officials about the potential rise of litigation based on past fiduciary decisions. That fear is heightened by the recent initiation of lawsuits by trial lawyers alleging some companies overcharged employees on fees for 401 (k) plans.

“There is a concern about potential lawsuits from employees alleging that a particular pension investment was imprudent,” agrees Jan. M. Jacobson, director, retirement policy, American Benefits Council, the main corporate pension lobby in Washington.

Lewis Freeman, president, Employers Council on Flexible Compensation, is one of many in the business community who want the DOL to bless capital preservation funds. “Such a vehicle may also be an appropriate default option for an employer with a very young population, or a high rate of turnover, where many of the plan's participants will terminate in short order and roll their account balances out of the plan,” he says.

But while support for inclusion of capital preservation products within the QDIA safe harbor is broad, only the life insurance industry wants the DOL to bless annuities. Ann B. Cammack, senior vice president, taxes & retirement security, American Council of Life Insurers, says, “The failure to include guaranteed insurance products, such as fixed annuity contracts, annuities with a fixed component, guaranteed investment contracts, stable value funds and other guaranteed products in the list of products eligible for QDIA status is an unacceptable shortcoming in the proposed regulation that must be addressed.”

But one financial industry executive, who did not want to be quoted, says that life insurance products are extraordinarily expensive. “I can’t imagine anyone would think they are an appropriate default investment,” she states.

Cammack responds, "Annuity products are a bargain when you consider what they offer, which is a guarantee of lifetime income. The fact is that you should not be building your retirement security on the cheap."

Not only are there disagreements about what kind of investments are appropriate for default 401(k) choices, but there is considerable unhappiness over the DOL’s preliminary decision to require a QDIA to either be managed by an investment manager or an investment company registered under the Investment Company Act of 1940. Schub says that requirement may greatly limit the ability of plan sponsors to offer independently-assembled “best in class” target-date or target-risk funds, which can be considerably cheaper than those offered by a mutual fund family.

The proposed rule also may limit the kinds of mutual funds which can be included in a QDIA. That is because it says a QDIA may not impose financial penalties or otherwise restrict the ability of a participant or beneficiary to transfer, in whole or in part, his or her investment alternative to any other investment alternative available under the plan. So would that exclude the many mutual funds who impose a redemption fee or a back-end sales load? That is the question Gail B. Mayland, vice president and associate general counsel, Charles Schwab & Co., Inc., is asking.

Given the lawsuits against Lockheed Martin, General Dynamics and some other companies on pension fund fees, companies are also nervous about what the DOL says about fees for automatic enrollment 401(k)s. The DOL proposed rule doesn’t address fees, except to rule out “financial penalties” when a participant transfers funds from one QDIA to another.

But the AARP, the lobby group for seniors, says that if QDIA fees are higher than for other comparable investments available on the market, then the plan fiduciaries must be able to justify choosing that investment for the QDIA. David Certner, legislative counsel and director of legislative policy for AARP, also wants DOL to publish fee disclosure guidance.

Health Insurers to Expand Offerings of Personal Health Records

December 2006 Digital Healthcare & Productivity.com

At a time when Nike's new Air Zoom Moire shoes send fitness data to a runner’s iPod Nano, the announcement on December 13 that health insurers were creating a portable, Web-based personal health record (PHR) was hardly revolutionary. In fact, speakers from the America’s Health Insurance Plans (AHIP) and Blue Cross and Blue Shield Association (BCBSA) at a press conference in Washington, D.C. used an infinite variety of rhetorical versions of the term “first step.”

The real significance of the announcement was as an impetus to the software industry to begin cranking up applications which could be used by consumers to maximize the value of these PHRs, and as a spur to convince physicians and hospitals to make a long-delayed start on ramping up office-based electronic health records systems which ultimately will be the prime beneficiary of these PHRs in a new era of real-time medicine.

The PHRs to be made available by AHIP and the Blues will cover 200 million individuals by the end of 2008. The data will be based primarily on claims received by the insurance company and consumer inputs on such things as immunization and family medical history. Scott Serota, CEO of the BCBSA, emphasized that the PHRs offered by individual companies will have tweaks beyond the core data elements, and will be “branded” for use as marketing tools. The PHRs depend for their portability on Health Level 7 and ANSI X12 protocols.

A plan member will be able to dictate what data is transferred from one health plan to another, or if that data should be provided to his or her physician. The data in the PHR will have all the privacy protections authorized by HIPAA and relevant state laws.

These PHRs are seen by the insurance industry as a way to help consumers improve their own health care, and as a way for the companies to cut costs associated with medical care that could otherwise be avoided. So a key component of these PHRs will be a constant sifting of medical claim, laboratory and pharmacy data against best practices and evidence-based guidelines, a process Aetna will do via what it calls its CareEngine. Aetna has actually offered that service, provided by a company called Active Health Management, to plan sponsors since 2002.

The challenge, of course, will be to get consumers to use these PHRs and, maybe more importantly, give physicians access to them, which is not technologically possible at the moment, given the low rates of electronic health record infrastructure adoption by the nation’s physicians and the absence of interoperability standards. As to the first challenge, AHIP and BCBSA have partnered with the National Health Council, which through its member groups has about 100 million members with various chronic illnesses. The NHC will be conducting pilot projects in an effort to educate its members on these PHRs, and stimulate their use.

The major benefit of the PHRs, however, is getting them into the hands of a patient’s physicians in real time, at the time of an examination, or when someone ends up in an emergency room. “But we are a ways away from creating an interoperable system,” acknowledged Bill Marino, CEO of Horizon Blue Cross Blue Shield of NJ.

OPENING THE DOOR TO FOLLOW ON PROTEINS

October 2006 issue of Biotechnology Healthcare

When the FDA approved the follow-on protein Omnitrope in May, it gave generic drug
makers the wedge they were hoping for. With pressure building in Washington, did
Omnitrope push the door open – or was it just an anomaly? BY STEPHEN BARLAS


When the U.S. Food and Drug Administration last May approved Sandoz’s Omnitrope — a follow-on protein to Pfizer’s Genotropin, the leading biotech human growth hormone
— hope sprung in the generic drug community that perhaps the agency had finally seen it their way, setting a precedent that would allow for the production and sale of follow-on proteins in the United States.
Omnitrope (somatropin) is the first follow-on protein from a generic pharmaceuticals company that the FDA has ever approved. Equally significant — and troubling
to the biotechnology indus-try — is the FDA’s near-withering 52-page reply to two biotech manufacturers and the Biotechnology Industry Organization, whose citizen
petitions had marshaled a phalanx of legal and regulatory arguments intended to persuade the
agency not to approve Omnitrope. Essentially, the three petitions said the FDA could not approve Omni-trope nor any other follow-on protein submitted via the 505(b)(2)
pathway, which otherwise permits a sponsor to rely on published studies or the agency’s finding of safety and effectiveness for an approved drug to support approval. The BIO
petition, for instance, cited significant differences between therapeutic protein products and chemical drugs, in terms of complexity and heterogeneity, and thus argued that the use of other companies’ data is no assurance of safety.
Perhaps more important than the approval of Omnitrope itself were the questions it raised. Was this a one-time shot, or did it create a defacto path for others to follow —even before a formal regulatory road map for follow-on biologics is developed and approved? Would
any future approvals be on a caseby-case basis? And how will Congress follow up?
Even though the FDA’s approval was epochal, the agency bent over backwards to refute that impression. In a Questions and Answers document posted on its Web site when it approved Omnitrope, the agency downplayed the precedent value of its approval by citing other
follow-on proteins it has approved under 505(b)(2): GlucaGen (glucagon recombinant for injection), Hylenex (hyaluronidase recombinant human), Hydase and Amphadase (hyaluronidase), and Fortical (calcitonin-salmon recombinant) nasal spray.
Tom Newton, PhD, pharmaceuticalmarket analyst for visiongain, a United Kingdom consulting company that recently published the exhaustive report Biogenerics 2006:
Challenges Ahead for an Emerging Market, says companies such as Novo Nordisk (GlucaGen), Halozyme (Hylenex) and PrimaPharm (Hydase) cannot really be thought of as “generic” manufacturers in the sense that Teva and Sandoz are. “In my opinion, that makes the application for Omnitrope significantly different from these examples,” he says. More importantly, he adds, all of the follow-on proteins alluded to by the FDA in its Q&A “appear to
be modifications of treatments already on the market, whereas Omnitrope is presented as a pure biogeneric.” Adds Newton, “This case has definitely raised the profile of biogenerics.
The authorities will come under increasing pressure to do something about it.”

WILL OTHERS FOLLOW?
Officials at top biotech companies concede that Omnitrope is likely to be succeeded by other
follow-on products, which is what has happened in Europe where follow-on proteins are referred to as “biosimilars.” David Beier, senior vice president for global government affairs at
Amgen, points to guidance documents published by the European Medicines Evaluation Agency
(EMEA). EMEA has developed both clinical and nonclinical guidances for recombinant products containing human insulin, somatropin(human growth hormone), granulocyte
colony stimulating factor (GCSF), and erythropoietin (EPO), and future guidance is expected on ainterferon and immunogenicity. Those publications, says Beier, led to the EMEA’s approval of Omnitrope and Valtropin, a recombinant human growth hormone, and its rejection
of a hepatitis C product. “Those European regulatory requirements in terms of safety, efficacy,
and pharmacokinetics are very, very similar to innovator products, but not identical,” Beier says. “In the main, although not in every detail, EMEA guidance has effectively
protected patient safety.”
Given the EMEA conditions for approval, and because in many cases newer innovative biotechnology medications are available that offer advantages over the older medicines that follow-on versions attempt to imitate, Beier feels that biologic follow-on products could
play a limited role in the marketplace by offering alternative products. But, he adds, the price advantage for biosimilars in Europe is substantially more modest than for the differential between brandname conventional drugs and generic copycat products. When they
first come on the market, European biosimilars offer a 10 to 20 percent price advantage over innovator drugs, according to Beier. Moreover, he says, there are fewer innovator
drugs coming off patent in the next five years than the generic industry “overestimates,” and biosimilars, which will have a less-robust safety profile, will offer no therapeutic advantage.
Ajaz Hussain, vice president and global head for biopharmaceutical development at Sandoz, agrees that the price differential in Europe for biosimilars will be smaller compared with traditional generic drugs. But, he argues, a follow-on product that costs that much less
than an innovator’s drug — whose annual cost to the patient may be in the $20,000 to $100,000 range —still amounts to an immense savings. Hussain declines to comment on how Sandoz will price Omnitrope in the United States.
Sandoz is already looking beyond Omnitrope, based on what Hussain describes as the FDA’s
precedent-setting decision, to other drugs the company could submit via the 505(b)(2) process. “We certainly plan to use it when we have a product that would fit the criteria,” he says, though he declines to be specific about possible candidates.

FDA: WHERE NEXT?
Omnitrope’s approval and the EMEA guidance documents would appear to point the FDA toward its next step: publication of guidance for approval of follow- on proteins submitted via
the 505(b)(2) pathway, which was developed before the emergence of complex biopharmaceuticals.
The U.S.agency has had some public workshops over the past two years, and has progressed on a guidance document in fits and starts, but it may be years before a guidance document is
drafted, vetted through public hearings, revised, and implemented. Established by the 1984 Drug Price Competition and Patent Term Restoration Act (commonly known as Hatch-Waxman), the 505(b)(2) pathway may be available for follow-on versions of drugs approved under section 505 of the Food, Drug, and Cosmetic Act, such as hGH and insulin, which are small-protein drugs containing few sugars —making them easier to duplicate.
Follow-on proteins in these categories are not exact copies, owing to the inexactitude of reproducing a drug via biotechnology — and, to be sure, Omnitrope itself is not rated as
therapeutically (AB) equivalent to any other human growth hormone and, therefore, is not substitutable for one. The 505(b)(2) pathway may be used, however, for a follow-on protein product that is sufficiently similar to an approved drug product to permit reliance, where scientifically justified, on certain existing information (including the FDA’s findings
of safety and effectiveness for an approved drug product) and may relieve the applicant from having to submit full-scale supporting data.
But GlucaGen, Hylenex, Hydase and Amphadase, and Fortical, all approved under 505(b)(2), are relatively simple molecules. The majority of biopharmaceuticals, infinitely more complex, are not approved as drugs under the Food, Drug, and Cosmetic Act, but licensed as biological products under section 351 of the Public Health ServiceAct — the “gold mine” route for
the generic drug industry. There is no approval pathway analogous to 505(b)(2) for products licensed under section 351.
That isn’t stopping generic drug makers such as Sicor, LG Chemicals, GeneMedix, Cangene, Rhein Biotech, Dr. Reddy’s Laboratories, Wockhardt, and Dragon Biotech from already supplying interferons, erythropoietin, and other biopharmaceutical products to Lithuania,
Mexico, China, Korea, India, Argentina, Egypt, Peru, and Brazil. These companies lick their chops at the prospect of getting FDA approval for those drugs, which are infinitely more complicated than human growth hormone and insulin.
Most observers, even those in the generic industry, agree that Congress would have to give the FDA new authority before it could approve generic versions of section 351 biopharmaceuticals. Amgen’s Beier notes that any new approval process for products regulated under section 351 must be constructed in a way that ensures patient safety and respects the intellectual property rights of the innovators. As Beier points out, a would-be sponsor of a followon biologic would be using a different cell line and different growth media to produce the
protein, and would likely use different fermentation methods, purification processes, and specifications. “Because of the inherent differences in these materials and processes, a generic sponsor cannot produce the same product as the pioneer,” he argues.
Genentech made a similar point in its April 2004 citizen petition, which asks the FDA to refrain from establishing standards for “similarity” of biotechnology-derived products under 505(b)(2). Alluding to safety concerns, the company maintained that “Current science
[does not] allow for reliance on analytical data and information generated from one biotechnologyderived product to support approval of a product manufactured through a different process.” Varied manufacturing processes, it pointed out, affect product purity
and can lead to immunogenicity. Genentech’s petition also gets to Beier’s point about respect for intellectual property rights, contending that any company that relies on another’s data to make safety and efficacy claims for a product that is not an exact copy of the innovator drug has, in essence, unfair access to the innovator’s trade secrets. The petition refers to a 505(b)(2) application from Dr. Reddy’s, an Indian company, for amlodipine maleate, based on the FDA’s approval of Pfizer’s amlodipine besylate (Norvasc), a calcium-channel blocker. The FDA stayed the effective date of the approval of Dr. Reddy’s product “because questions [were] raised
about the source of the data the [FDA] relied on in approving” the application, according to the petition. “We are similarly concerned about the protection of our confidential commercial information.”
Congress is notably slow-footed, so whether legislation pertaining to section 351 passes in the near future is unlikely. What is more likely over the next six months to a year is the submittal of additional 505(b)(2) applications by major generic drug companies.

FIGHTING BACK
Of course, Pfizer is worried about Omnitrope. In its May 2004 citizen petition, Pfizer argued that the FDA could not legally approve Omnitrope via 505(b)(2) because the agency would have to access confidential Pfizer manufacturing and clinical data to do so. The Genentech and BIO petitions made a different point: The FDA could not issue guidance for generic drug companies on how to navigate the 505(b)(2) process, though they cited reasons similar to the ones Pfizer
used in its anti-Omnitrope petition.
When it approved Omnitrope, the FDA sent a 52-page explanation to each of the three petitioners, carefully knocking down every one of the industry’s objections like pins at the end of a bowling lane — and seemingly with enough force that those pins could fly across lanes.
Briefly, the agency said that the active ingredient in Omnitrope, somatropin, is highly similar to the
active ingredient somatropin in
Genotropin. Sandoz was able to
demonstrate that Omnitrope was
“sufficiently similar” to Genotropin
to warrant reliance on FDA’s finding
of safety and effectiveness for
that drug. Somatropin is the active
ingredient that has been part of
seven different recombinant hGHs
the agency has approved since somatrem
(Protropin) in 1985. Sandoz
also provided extensive independent
evidence of Omnitrope’s
safety and effectiveness for use in
pediatric patients with growth hormone
deficiency through three sequential,
multicenter, phase 3 pivotal
trials over a 15-month period,
along with other supportive data.
Moreover, the FDA said it did not
depend on any trade-secret information
submitted by Pfizer to approve
Omnitrope.
So the FDA seemed to be arguing
that small follow-on proteins, as
opposed to big molecules — at
least in a couple of therapeutic categories
— could be approved without
reliance on brand-name trade
data, and the FDA will continue to
do just that. The unstated message
was: stop complaining and live
with it.
Whether Pfizer accepts that message
is another matter. Paul Fitzhenry,
a spokesman for Pfizer, says
his company has not yet decided
whether to take the FDA to court
over its approval of Omnitrope.
The Genentech and BIO petitions
have even broader implications, because
they argued that the FDA does
not have the authority to either approve
follow-on proteins or guidance
for generic drug companies.
Walter Moore, vice president of government
affairs for Genentech, says
his company has made no decision
yet on whether to take the FDA to
court. But he adds, “We do not consider
the FDA letter a full response to
our citizen petition by any means.”
Does the FDA even need to issue
guidance at this point? Its answer
to the petitions may be guidance
enough for companies making follow-
on protein products that fit an
Omnitrope-type profile. An FDA
spokesperson, Karen Mahoney says
that, “The Agency will not comment
on the status of unpublished
draft guidance.”
WHAT WILL CONGRESS DO?
Some members of Congress believe
that FDA guidance on how
follow-on protein products can
clear through 505(b)(2) would be a
useful political exclamation point,
such as Republican Sen. Orrin
Hatch of Utah and California Democratic
Rep. Henry Waxman,
authors of the landmark 1984 law
that opened the door to expedited
FDA approval of abbreviated applications
from generic drug companies.
Earlier this year, Hatch and Waxman sent a letter to the FDA
asking it to publish guidance on
hGH and insulin, two categories
amenable to follow-on proteins.
On Sept. 29, Waxman, along with
New York Democratic Sen. Charles
Schumer, introduced the “Access to
Life-Saving Medicines Act,” which
would amend section 351 to approve
abbreviated applications for
biologics that are “comparable” to
previously approved products. The
bill defines comparable as demonstrating
no clinically meaningful
differences. Introduced a
week before Congress recessed
for elections, the bill got its
sponsors some publicity back
home but won’t see any action
before the the 109th Congress
officially closes next month.
A more likely congressional
response to the FDA’s consideration
of Omnitrope would
be legislation forcing the FDA
to make decisions promptly on
individual 505(b)(2) applications,
even when citizen petitions
are filed. Innovator biotech
companies have filed a couple dozen
citizen petitions in the past few years
in an effort to prevent FDA approval
of both follow-on proteins and conventional
generics.
At hearings the Senate Select
Committee on Aging on July 20,
Gary Buehler, director of the FDA’s
Office of Generic Drugs, said that
an agency evaluation of 42 petitions
answered between 2001 and 2005
showed that 33 had been denied in
full, 3 denied in part, and 6 granted.
“While the citizen petition process
is a valuable mechanism for the
agency to receive information from
the public, it is noteworthy that very
few of these petitions on generic
drug matters have presented datatherapeutically (AB) equivalent to
any other human growth hormone
and, therefore, is not substitutable
for one. The 505(b)(2) pathway may
be used, however, for a follow-on
protein product that is sufficiently
similar to an approved drug product
to permit reliance, where scientifically
justified, on certain existing information
(including the FDA’s findings
of safety and effectiveness for an
approved drug product) and may relieve
the applicant from having to
submit full-scale supporting data.
But GlucaGen, Hylenex, Hydase
and Amphadase, and Fortical, allapproved under 505(b)(2), are relatively
simple molecules. The majority
of biopharmaceuticals, infinitely
more complex, are not
approved as drugs under the Food,
Drug, and Cosmetic Act, but licensed
as biological products under
section 351 of the Public Health Service
Act — the “gold mine” route for
the generic drug industry. There is
no approval pathway analogous to
505(b)(2) for products licensed
under section 351.
That isn’t stopping generic drug
makers such as Sicor, LG Chemicals,
GeneMedix, Cangene, Rhein
Biotech, Dr. Reddy’s Laboratories,
Wockhardt, and Dragon Biotech
Pressure is likely to mount on
Congress and the FDA to address
the absence of section 351 followon
protein products because of the
cost of drugs like Epogen, Procrit
and Eprex — the three biggest selling
versions of epoetin alpha, a recombinant
form of erythropoietin,
which is a hormone that stimulates
the production of red blood cells.
Epoetin alpha brands and their derivatives
are the most successful
biotech drugs on the pharmaceutical
market. The first process
patent to expire for epoetin
alpha lapsed in 2001 in Europe
and 2004 in the U.S., according
to visiongain. Most epoetin
products are already off patent
— approximately 70 percent
of the total market has lost
patent protection.
Former Deputy FDA Commissioner
William Schultz,
now a Washington lawyer
who has represented the
Generic Pharmaceutical Association
and generic companies,
says a coalition of business and
health groups is pushing Congress
to give the FDA the authority it
needs to approve more follow-on
protein products. “Cost of biopharmaceuticals
is pushing this issue,”
says Schultz, “not just from generic
companies and consumers, but
from payers in the private sector
and the federal government.”
Congress may get its first chance
to amend section 351 in 2007 when
Congress must reauthorize the Prescription
Drug Users Fee Act. It will
be an event worth watching. BH
Stephen Barlas has covered the FDA and
drug issues since 1981 when he became
a full-time freelance Washington journalist
for business and trade publications.

Washington Letter: FMEA as a Packaging Tool

September 2006 issue of Pharmaceutical Manufacturing

By Stephen Barlas, Washington Correspondent

A July report on medication errors by the Institute of Medicine (IOM) underscored the utility of Failure Modes and Effects Analysis (FMEA) in managing risk. The FDA already endorses FMEA, describing it in a June Quality Risk Management guidance document as “powerful tool.” In fact, the FDA even uses FMEA itself, to help weed out confusing drug names once a new drug application (NDA) has been submitted.

But neither the FDA nor the drug industry use FMEA to assure the development of clear, consistent drug labeling and packaging. According to the IOM’s latest report, failure to use FMEA leads to such problems as:

  • cluttered labeling
  • small font
  • serif typeface
  • lack of background contrast
  • inadequate prominence of reminders and warnings
  • overemphasis on company logos and trade dress

— all of which “continue to have a direct effect on the readability and comprehensibility of product labels, and hence on rates of medication errors.”

The report suggests that the FDA require FMEA analysis for all pharmaceutical labeling and packaging design and assessment, and that drug manufacturers be required to submit those assessments as part of any new drug application (NDA). It also urges the FDA to publish two separate guidance documents, one on naming, the other on labeling and packaging, by the end of 2006, and to encourage industry to expand unit-of-use packaging to new therapeutic areas.

The Pharmaceutical Research and Manufacturers of America has taken no official position on whether FMEA should be required, according to PhRMA associate vice president Alan Goldhammer. In an unsigned response to the IOM report, the FDA alluded to guidance documents on naming, labeling and packaging that it is planning for later this year. So far, the FDA has not weighed in on the IOM report recommendations.

Progress Being Made on Medication Errors

The IOM report asserts that some progress has been made toward reducing medication errors since an earlier advisory committee issued a report on the topic six years ago. However, there is still plenty of room for improvement. “The frequency of medication errors and preventable adverse drug events is cause for serious concern,” said committee co-chair Linda R. Cronenwett, dean and professor, School of Nursing, University of North Carolina, Chapel Hill.

Sen. Charles Grassley (R-Iowa), chairman of the Senate Finance Committee and a key congressional Medicare decision maker, issued a statement after the report’s release highlighting the IOM recommendation that the FDA issue guidance on drug naming, labeling and packaging by the end of 2006.

With regard to the recommendation on wider implementation of unit-of-use packaging, drug companies have been moving in that direction. This is due, in large part, to an FDA requirement that kicked in last April, which states that all drugs going to hospital pharmacies must have a linear bar code containing, at a minimum, the drug’s National Drug Code (NDC) number.

The FDA rule does not require that hospital SKUs (stock keeping units) be packaged in a unit-of-use or unit-dose package. But Reynard Jackson, executive vice president, business development, packaging services at Cardinal Health, says that most of his company’s 200 pharmaceutical packaging clients are beginning to do just that. They’re complying, he says, with the “spirit” of the FDA rule, which is aimed at reducing medication errors. The FDA just approved two Pfizer hospital blister packs for Lipitor (atorvastatin calcium), for example.

“The question is,” asks Opal Johnson, marketing director at Pearson Medical Technologies, “how fast are they [drug manufacturers] moving on this? What are there — 80,000 drugs out there?” Pearson sells its intelliPack2 blister packaging and m:Print bar code labeling equipment to smaller hospitals.


streetwise

From September 2006 Strategic Finance

S T E P H E N B A R L A S , E D I T O R

Section 404 Saga Continues

The Securities & Exchange Commission will issue guidance for corporate
management on how to comply with the provisions of Section 404 of the
Sarbanes-Oxley Act, which requires companies to explain their internal controls
and test them. The SEC issued a “Concept Release” in mid-July that outlined
some of the areas it may address when the guidance is published. No
date for publication was mentioned. Large companies, of course, have had to
comply with 404 for two years. Smaller companies received an initial reprieve,
but, with first-time compliance looming, they are pushing the SEC hard to
make some changes in required compliance. The SEC Advisory Committee on
Smaller Public Companies raised a number of concerns in an April report on
the ability of smaller companies to comply with 404 in a cost-effective manner. That was followed days later by a U.S. Government Accountability Office report that said much the same thing. Of course, larger companies and the U.S. Chamber of Commerce have been arguing for
the past year that Section 404 is too costly. The SEC took a first crack at quieting that murmuring when it issued limited guidance in April 2005 relating to the exercise of professional judgment, the concept of reasonable assurance, and the permitted communications between management and auditors. But this next round of guidance will go further. For example, the 2005 guidance stated that management needs to use “reasoned judgment” in identifying internal controls. But few understood what that meant. The July 2006 Concept Release maintains that “many companies did not efficiently and effectively identify risks to reliable financial reporting and relevant internal control functions, ultimately leading to the identification, documentation, and testing of an excessive number of controls.We are also
skeptical of the large number of internal controls that some companies have
identified, documented, and tested.” Part of the reason for that problem is that corporate management, in implementing Section 404, may have leaned too heavily on the Public Company
Accounting Oversight Board (PCAOB) Auditing Standard No. 2 (AS2), An Audit of Internal Control over Financial Reporting Performed in Conjunction with an Audit of the Financial Statements. Published in June 2004, this was meant for outside auditors, not management. But many corporate financial and accounting departments interpreted AS2 very conservatively, increasing costs far beyond what was necessary. This is

c o n t i nue d o n p a ge 2 4

the problem that the SEC hopes to resolve with its new guidance, which the agency emphasized will be “scalable.” That apparently means the guidance won’t be written only with
small companies in mind but also will take the concerns of Fortune 500 companies into account.

New Chief Accountant

With more expansive 404 guidance now in the SEC pipeline, Conrad Hewitt, the SEC’s new chief accountant, joins the Commission at a critical time. The post was vacant since
Donald Nicolaisen departed last fall. Hewitt, who had been serving on three corporate board audit committees, had otherwise been retired from a full-time job since 1998,
when he was a California state banking official. From 1972 to 1995, Hewitt was the managing partner of Ernst & Young and its predecessor firm, Ernst & Ernst, in the firm’s
Northern California (1986-1995), Seattle (1979-1986), and Honolulu (1972-1979) regions. In his statement upon joining the SEC, Hewitt noted that he has worked with many of the big accounting firms to implement Sarbanes-Oxley and looks forward to working with the PCAOB
“to maximize the protection of shareholders while eliminating excessive costs and burdens both
here and abroad.”

Bill Would Eliminate Some State Taxation of Corporations

Protests from states and local governments forced House Republican leaders to postpone a floor vote on a bill limiting state taxation of corporations. A spokesman for Rep. John Boehner (R.-Ill.), the House Majority Leader, said, “Some misperceptions about [the bill’s] effect on
states” resulted in the vote being delayed, probably until after the August recess. The bill (H.R. 1956) would establish a national standard for when states can collect business activity taxes from multistate companies whose principal locations are outside the state but who have many
customers within the state. The bill has been strongly supported by companies in the retail, financial, and medical industries. The bill would require companies to have a physical presence in a state for at least 21 days before they could be taxed. But there would be some exceptions. The National Governors Association spooked both Republicans and Democrats in the days prior to the scheduled House floor vote by publicizing a study that projected revenue losses to the states would be nearly double the annual losses of $3 billion by 2011 that the Congressional
Budget Office projected. Besides business activity taxes, states couldn’t levy gross receipts, license, or franchise taxes.

Stricter U.S. Gas Standards Stalled


IEEE Spectrum from September 2006

By Stephen Barlas
Bipartisan opposition defeats bipartisan efforts to strengthen CAFE regulations

Gasoline may be more expensive than ever in the United States, but talk is still cheap on Capitol Hill. Flocks of bills promoting automobile fuel efficiency and alternative fuels, which took flight on soaring rhetoric last spring, have dropped to the ground like so many downed birds. Nearby, the special interests stand with shotguns still smoking.

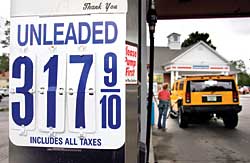
Congress had tried in previous sessions to increase Corporate Average Fuel Economy (CAFE) standards for passenger automobiles from a fleetwide average of 27.5 miles per gallon (8.55 liters per 100 kilometers), where they have been stuck since 1985. With gas prices at more than US $3 per gallon [see photo, “High Prices, Big Cars”], Representatives Sherwood Boehlert (R‑N.Y.) and Edward Markey (D-Mass.) thought that they had the political impetus they needed to pass a bill that would increase CAFE standards for both cars and light trucks—light trucks have had a separate standard—to 33 mpg (7.13 L/100 km).

Boehlert, who will retire this year as chairman of the House Science Committee, has been tirelessly pounding home the main message of the influential 2002 report issued by the National Research Council, Effectiveness and Impact of Corporate Average Fuel Econ­omy Standards. “The technologies needed to meet the standards our bill sets already exist,” Boehlert says. “Indeed, some of them have already been surpassed since the report was issued in 2002.”

Rather than boost CAFE standards as such, the Bush administration has preferred to give the National Highway Traffic Safety Administration the authority to change the way the standards are figured for passenger cars to achieve some of the same effect. It has moved to a method that focuses on footprint, a measure of a car’s wheelbase, and away from automobile weight and fleet averages [see box, “Calculating CAFE”].

Last March the highway safety administration, which already had the authority to use the footprint method to determine CAFE standards for the light truck category, adopted that method for cars as well. The result is that average miles per gallon for sport utility vehicles (SUVs), small trucks, and minivans will increase from 21.6 mpg on 2006 models to 24 mpg for 2011 models on an industrywide scale. The increase is better than it looks, because huge vehicles like Hummers were included in that group for the first time.

But when a footprint bill came be­fore the House Energy and Commerce Com­mittee in May and passed, Markey tried to attach an amendment with his preferred fuel mileage boost to 33 mpg. He lost by a vote of 36–17. Michigan Representative John Dingell, the committee’s senior Democrat and a close ally of the automobile industry, led a number of other Democrats in opposition to the Markey amendment, which was also opposed by some Republicans.

In the estimation of Eli Hopson, the Washington representative of the Union of Concerned Scientists, not even the underlying footprint bill, which made it through the committee, will pass the full House. That view is seconded by Joe Pouliot, Boehlert’s spokesman, who says there are three separate camps of House members with three mutually exclusive approaches to higher fuel efficiency, none of which commands a House majority: (1) no CAFE changes, no way; (2) CAFE light, that is to say, the footprint approach; and (3) CAFE heavy, the Boehlert-Markey alternative.

Inaction suits the auto companies just fine, and this goes as well for the main union representing their employees, the United Auto Workers. Alan Reuther, the legislative director of the UAW, told members of the House committee that imposition of a higher miles-per-gallon requirement “would severely discriminate against full line [auto] producers whose product mixes contain greater percentages of larger cars and light trucks.” The UAW also opposes a shift to a footprint calculation for autos, because, it says, dropping the fleetwide average would theoretically allow the Big Three U.S. automakers to leave the production of smaller cars to foreign manufacturers. In the current system, U.S. carmakers need to make small cars in the United States to balance out their many SUVs; a footprint system would render that balancing unnecessary.

In the Senate, Democrat Dianne Fein­stein of California and Republican Olym­pia Snowe of Maine have been pushing a 35-mpg (6.72–L/100 km) bill. But some senators who normally are attuned to the environment and concerned about the country’s ever greater dependence on foreign oil have been even more sensitive to the concerns of autoworkers. For example, New York Senator Hillary Rodham Clinton, also a Democrat, has gently spoken in favor of boosting fuel economy “responsibly without needlessly sacrificing safety or American jobs.” Meanwhile, fuel economy standards are much stronger in some other places, from Europe to China [see graph, “U.S. Fuel Standards Lag ”].

The auto industry has argued that it is too expensive, for both manufacturers and consumers, to adopt the engine and transmission technologies identified in the 2002 National Research Council report. Instead, they say that the way to conquer the United States’ addiction to foreign oil is to roll out more alternative-fuel vehicles, a vision the companies are propounding in print ads and Washington-read opinion magazines. The auto companies point out that there are already 8 million alternative vehicles on the road in the United States and that there will be 1 million more by the end of 2006.

Of that total, boast the companies, there are 21 different car makes—5 ­million vehicles in all—that run on E85, a mixture of 85 percent ethanol and 15 percent gasoline. But E85 is more expensive than ­gasoline, it provides inferior fuel efficiency, and it yields little if any reduction in greenhouse gas emissions. What’s more, “there is currently little customer demand” for E85 vehicles, concedes Reg Modlin, director of environmental and energy planning for DaimlerChrysler Corp., and there are only 650 service stations nationwide that carry E85. “Congress should help in accelerating the growth of ethanol production, distribution, and retail-sales infrastructures through tax incentives, capital depreciation allowances, or other fiscal instruments,” Modlin says.

Sponsorship of E85 infrastructure legislation has already begun: a couple of bipartisan teams have cropped up in Congress, paralleling the Boehlert-Markey partnership. The Biofuels Secur­ity Act, for example, would require major oil companies to increase the number of E85 pumps at their service stations by five percentage points a year.

In addition, the biofuels bill would create a new consumer tax credit for the purchase of “flexfuel” vehicles if the vehicles have no fuel efficiency loss from the use of E85 as compared to regular gasoline. “We understand that there is technology available—for example, a Saab flexfuel E85 vehicle on the market in parts of Europe—allowing vehicles to have no fuel efficiency loss when burning E85 in comparison to gasoline, and perhaps even some mileage gain,” says Senator Tom Harkin, an Iowa Democrat and a cosponsor of the bill.

Disputing that view is Susan Cischke, vice president of environmental and safety engineering at Ford Motor Co.: “We’ve heard from many people that all it takes to make a flexfuel vehicle is ‘a little tweak to the chip that runs the engine.’ I wish it were that simple, but it’s not.”

Big oil has big problems with the biofuels bill, mainly because of its five-percentage-point mandate, and with other ethanol bills, such as the “10 by 10 Act,” which would require refiners to blend at least 10 percent ethanol into each gallon of gasoline by 2010. Autos can use a 10-to-90 blend without any changes to an engine.

At the American Petroleum Institute, an organization in Washington, D.C., that represents the big oil companies, Ed Murphy, group director of refining marketing, says that the institute strongly opposes any bill that mandates installation of E85 pumps and tanks. He argues that even if 100 percent of service stations offered E85, consumers wouldn’t buy it because of its higher cost and lower performance. E85 proponents, Murphy complains, are engaged in “a cynical attempt to appear green.”

It appears, in any case, that the institute has little to worry about. Bob Dinneen, president of the Renewable Fuels Association, an ethanol industry group in Washington, D.C., says he has little hope, realistically, that Congress will enact any kind of biofuels legislation any time soon. “It is a fairly dysfunctional Congress right now,” he observes glumly.

WHOLE GRAIN LABELING

From August 2006 issue of Packaging World magazine


FDA draft guidance ignites debate on package claims regarding the use of whole-grain claims.

Stephen Barlas, Contributing Editor

The Food and Drug Administration’s attempt to referee the whole-grain foods labeling debate has various food companies crying foul.

General Mills, Inc., and a number of allies are using the draft guidance the FDA issued last February as the latest basis for criticizing the agency for refusing to establish a safe harbor for the use of terms like “good” and “excellent” source of whole grains.

General Mills submitted a petition to the agency in 2004 asking to be able to label its cereals and other products as good and excellent sources of whole grains if they met a certain content standard. The FDA never acted on that petition, which didn’t bother the company. In late 2004, GM went ahead anyway and reformulated its breakfast cereals in order to be able to label them as “good” and “excellent” sources of whole grains based on the criterion used by the Whole Grains Council (WGC), a private group made up of various industry associations.

“Cereal companies and baked goods companies are arguing over when a company can use the term ‘whole grain’ on its label.”

To date, the FDA has not defined, in either a guidance document or law, how much whole grain has to be in a serving of a product for the company to be able to use the description “good” or “excellent” source of whole grain on the package. In the draft guidance it issued in February, the agency signaled it intended to stick to that position.

Nonetheless, it isn’t technically illegal to use those terms. The FDA has no law against such descriptive terms on food labels. But neither is use of the label claim exactly kosher, as it would be if the FDA approved guidance on that subject. Doing so would create a “safe harbor” for companies, meaning there would be no chance of FDA enforcement action against them as long as they heeded the terms of the guidance.

In this current grey area environment, companies like General Mills who use those terms run the risk, however slight, of the FDA taking enforcement action against them.

Push for protection

That explains the push by General Mills, the Whole Grains Council and their allies to convince the FDA to change its mind. The WGC has offered members, including GM, use of a stamp design proclaiming a product a “good,” “excellent,” or “100% excellent” source of whole grains.

The stamp was made available in January 2005, just after the U.S. Department of Agriculture published its Dietary Guidelines for Americans, 2005 that recommended that individuals “consume 3 or more ounce-equivalents (48 grams) of whole-grain products per day, with the rest of the recommended grains coming from enriched or whole-grain products.”

“To date, the FDA has not defined, in either a guidance document or law, how much whole grain has to be in a serving of a product for the company to be able to use the description ‘good’ or ‘excellent’ source of whole grain on the package.”

The WGC established 8 grams of whole grains as the amount a product must contain to use a “good” label, 16 grams for “excellent,” and the product must be 100% whole grain to use the “100% excellent” label.

That stamp has proven very popular, providing some evidence that whole-grain labeling of food is becoming the phenomenon that perhaps “low-carb” once was. There were 27 members in the WGC back in January 2005. Today, there are 116, according to Cynthia Harriman, director of food and nutrition strategies.

General Mills uses the stamp on its Cascadian Farm granola bars, although not on its breakfast cereals, many of which do have “good” or “excellent source” label copy on their packages. A few of the other major companies using the whole grain stamp are Kashi (Kelloggs) and American Italian Pasta Co (pasta—Muellers noodles).

Guidance worries

The FDA’s announcement in the draft guidance that it continues to oppose use of “good” and “excellent” source of ingredient claims for whole grains applied new pressure to food manufacturers. It led to a meeting between WGC and FDA officials on March 30.

Subsequently, in June, the WGC announced that it was changing its stamp by requiring users to include language stating either the actual number of whole grain grams in a serving, or the fact that a product has “more than” eight or 16 grams of whole grain. Each of the stamps will also say, “Eat 48g or More of Whole Grains Daily.” General Mills, for example, which doesn’t use the stamp, but does make “good” source claims on its cereal packages, is also adding that quantitative copy to its packages, according to spokeswoman Kirstie Foster.

However, even with the addition of the quantity of grams and the USDA recommendations, the WGC stamp would still seem to run afoul of the guidance provided in the February draft document. That is just fine with Campbell Soup Co., the leading opponent of an ingredient content claim for whole grains. Campbell opposed General Mills’ petition in 2004, and has been dogging the cereal giant’s heels ever since. Chor San Khoo, vice president global nutrition & health at Campbell, says, “Campbell supports the FDA in opposing statements that characterize a particular level of whole grains as a good or excellent source.” Campbell argues that whole grains are not a nutrient, so no Daily Value, which is a USDA term of art, exists.

Other concerns

While the debate over ingredient source claims for whole grains is the most controversial issue arising from the draft guidance, it isn’t the only one.

More broadly, cereal companies and baked goods companies are arguing over when a company can use the term “whole grain” on its label. In the draft guidance, the FDA states: “Depending on the context in which a “whole grain” statement appears on the label, it could be construed as meaning that the product is ‘100% whole grain.’ We recommend that products labeled with ‘100% whole grain’ not contain grain ingredients other than those the agency considers to be whole grains.”

“Depending on the context in which a ‘whole grain’ statement appears on the label, it could be construed as meaning that the product is ‘100% whole grain’.”

This doesn’t sit well with manufacturers of frozen pizzas and breads. “Requiring pizza crust to be made with 100% whole-grain flour in order to define a whole-grain pizza will likely delay, if not block, pizza incorporation of significant amounts of whole grain into products,” states Bruce Paterson, vice president of research and development for The Schwan Food Co., which sells Freschetta and Red Baron frozen pizzas, and other frozen foods.

William Matthaei, president of Roman Meal Co., which uses the tagline, “Natural Whole-Grain Goodness,” says the requirement that breads be 100% whole grain to make that claim on the label would pose a problem for his products. Those products employ vital wheat gluten, a concentrated wheat protein that does not contain any of the wheat bran or germ. And those are identified in the draft guidance as part of a whole grain.